Abstract
There are many approaches to molecular energy calculations, and there are still no methods to calculate the binding energy of a molecule accurately. Also, in considering molecules, all processes are actually performed at finite temperatures. And the process of molecular formation is a process of increasing the binding energy of a molecule, and if the molecule is considered as a thermodynamic system, it is also possible to think of the physical quantity that determines the direction of the spontaneous process. This paper propose a new approach to calculating the binding energy of a molecule by examining the relationship between the electron density and binding energy and about the thermodynamic formalization of molecule. From the calculated value by Gaussian09w (HF, 6-31g) simulation of 100 molecules of 10 species and the binding energy in some literatures, the correlation was derived and confirmed that two parameters are given for each species. It was found that the binding energy of a water molecule can be calculated, in particular. Also the molecular free energy is conceptualized where the value can be index of molecular formation and molecular optimization process. This theory, together with the potential well theory in quantum mechanics, can serve as a basis for explaining the phenomenon that for molecules with the same chemical structure formula, higher energy molecules exist more stably.
Keywords
Thermodynamic Formalization, Electron Density, Binding Energy
1. Summary
To calculate molecular energy, atomic charge, and molecular structure, wave function theory (WFT)
| [1] | E. Schrödinger, An Undulatory Theory of the Mechanics of Atoms and Molecules. Phys. Rev. 28, 1049–1070 (1926). |
| [2] | Hartree, D. R. The Calculation of Atomic Structures. Rep. Prog. Phys. 11, 113-43 (1947). |
| [3] | Kohn, W. Nobel Lecture: Electronic Structure of Matter-Wave Functions and Density Functionals. Rev. Mod. Phys. 71, 1253–1266 (1999). |
| [4] | Miyajima, K., Yabushita, S., Knickelbein, M. B. & Nakajima, A. Stern-Gerlach Experiments of One-Dimensional Metal-Benzene Sandwich Clusters: Mn(C6H6)m (M) Al, Sc, Ti, and V). J. Am. Chem. Soc. 129, 8473–8480 (2007). |
| [5] | DiBenedetto, S. A. et al. Structure-Performance Correlations in Vapor Phase Deposited Self-Assembled Nanodielectrics for Organic Field-Effect Transistors. J. Am. Chem. Soc. 131, 11080–11090 (2009). |
| [6] | Prasad, V. K., Otero-de-la-Roza, A. & DiLabio, G. A. Atom-Centered Potentials with Dispersion-Corrected Minimal-Basis-Set Hartree-Fock: An Efficient and Accurate Computational Approach for Large Molecular Systems. J. Chem. Theory Comput. 14, 726–738 (2018). |
[1-6]
and density function theory (DFT)
| [3] | Kohn, W. Nobel Lecture: Electronic Structure of Matter-Wave Functions and Density Functionals. Rev. Mod. Phys. 71, 1253–1266 (1999). |
| [7] | Kohn, W. & Sham, L. J. Self-Consistent Equations Including Exchange and Correlation Effects. Phys. Rev. 140, A1133–A1138 (1965). |
| [8] | Slater, J. C. Quantum Theory of Matter, 2nd ed.; McGraw-Hill: New York, Chapter 16. 68). |
[3, 7, 8]
are proposed, and several methods (ab initio, semi-empirical methods so on) has been established, so the calculation for many projects are processed and recently, the pseudo chemical potential (PCP) theory
| [9] | T. I. Kim et al, A Novel Method for Calculation of Molecular Energies and Charge Distributions by Thermodynamic Formalization, Scientific Reports 9, (2019) 20264. |
| [10] | Jon Yung et al, Study on Calculation Method of Parameters in Molecular Calculation Model by Pseudo Chemical Potential Method, Science Research, (2023) 11(3): 43-50. |
| [11] | Jong Yong Ju et al, Theoretical Base of Pseudo Chemical Potential (PCP) Method, Science Research, (2022) 10(6): 137-143. |
[9-11]
proposed from thermodynamic formalization of molecule and reported higher accuracy and faster calculation.
Wave function theory has been used for molecules with high computational accuracy but low atomic number due to high computational effort, and density functional theory has been used to calculate molecules with high atomic number. Although electron density theory has improved both computational and computational speed, it is still computationally expensive for macromolecules.
Hence, our workgroup has previously proposed the PCP theory by thermodynamic formulation of molecules and presented the data for calculation of total energy of molecules
| [9] | T. I. Kim et al, A Novel Method for Calculation of Molecular Energies and Charge Distributions by Thermodynamic Formalization, Scientific Reports 9, (2019) 20264. |
[9]
. The above methods calculate the total energy of a molecule with high accuracy, but the calculation of the binding energy of a molecule does not work well. The aim of this paper is to present a method for calculating the binding energy of a molecule and to elucidate its theoretical basis and effectiveness.
Molecular energy can be viewed as the sum of the atomic region energy of a molecule and the interaction energy (binding energy) between the atomic region.
(1)
The value of the electron density of a selected atom in a molecule can be determined by the influence of all atoms in the molecule, which means that the value of the electron density can reflect the energy of the interaction of atoms in a comprehensive way. The binding energy can be calculated as a function of the electron density as a variable.
where is a function of the total electron density of atoms in a molecule as a variable, and E0 is a quantity with energy content.
According to the PCP theory, the molecule is formulated as a thermodynamic system consisting of a multiphase one component, and thermodynamic laws are also established in the molecule
| [11] | Jong Yong Ju et al, Theoretical Base of Pseudo Chemical Potential (PCP) Method, Science Research, (2022) 10(6): 137-143. |
[11]
.
(3)
where θ is temperature of the thermal equilibrium system, A is the free energy of θ, 𝜈(r) system, E is the total energy of system and S is entropy of electron system under θ, 𝜈(r).
This leads to the idea that molecular thermodynamic systems can also consider the free energy of a molecule and can be a measure of the direction of the molecular formation and molecular optimization process.
2. Theoretical Foundation
2.1. Relationship Between Electron Density and Binding Energy
The electron density is density function of electron with respect to the position in the multi-electron system. The electron density is given by (ψ- wave function, ψ*- complex conjugated wave function, d-volume element) and can be found by molecular orbital and atomic orbital methods, respectively.
In the molecular orbital method (LCAO), the electron density ⍴r of the r atom is defined by the following expression:
where νi is the number of electrons in the i-molecule orbital and Cir is the coefficient of the atomic orbital function of the r-atom in the i-molecule orbital.
The larger Cir is, the higher is the probability that the electron in the i-molecule orbital is around the r atom. This is the electron density in the r atom of the electron in the i-molecule orbital. This is the total electron density of the r atom when it is added to all the electrons of all orbital.
The PCP theory based on thermodynamic formalism has formulated molecules as systems in thermodynamic systems, multi-phase systems with atoms as phases and electrons as components.
The formation of molecules occurs at finite temperature, not at zero temperature limit. Therefore, it is also important to consider the entropy change of this process in the thermodynamic formulation of the molecular system. From the viewpoint of thermodynamics, the spontaneous process in a molecular system is a process in which the free energy of the molecular system decreases at constant temperature and volume, and reaches a minimum at equilibrium. That is
For an infinitesimal equilibrium process of a molecular system under isothermal conditions, the free energy change is expressed as
where A is the free energy of the molecular system, E is the internal energy, is the temperature, and S is the entropy.
When the corresponding thermodynamic quantities for a molecular system in dissociation at the same temperature are expressed as A0, E0, and S0, respectively, these quantities are constant, so we can rewrite the above equation as follows.
Here
This is the difference between the thermodynamic quantities of the molecular system for the stable and dissociated states of the molecule. E is the binding energy of a molecule in physical sense.
The above equation shows a linear relationship between ∆E and ∆S in the equilibrium process of molecular system change. That is
This equation allows us to relate θS to the binding energy at equilibrium.
Here
Since the electron density in a molecule changes when a molecule goes from a dissociative state to a stable molecular state, the entropy change of this process can be related to the electron density.
This allows us to guess the following functional relationship between the binding energy and the electron density:
Introducing this relation, A is expressed in equilibrium as follows.
Consequently, it may be more realistic to minimize the free energy A of this system at finite temperature instead of minimizing the E of the molecular system in the absolute temperature zero limit.
We explored the meaning of entropy to concrete the relationship between entropy change and electron density.
In a thermodynamic system, entropy is a quantity that characterizes the number of microscopic states that belong to the system. The number of states in a molecule is determined by the probability that an electron can appear, expressed as the electron density.
Also, entropy is a value that indicates disorder. The electron density, which represents the probability that an electron can appear around an atom in a molecule, can also be considered to represent the disorder of electrons. In thermodynamic systems, the entropy value is small if the components are arranged separately, and the entropy value is large if they are uniformly distributed. When the electron arrangement (the probability of appearing around an atom) is biased by a highly polar atom in a molecule, the electron density of atoms increases as the number of electrons in the cavity region overlapping the atomic regions increases, and decreases when the arrangement of electrons is uniform.
In the above, it is conceivable that in a molecular system, we should consider the electron density instead of the entropy in existing thermodynamic system. And the electron density is a value that reduces the probability that electrons appear around an atom, so the relationship between the electron density and the binding energy should be calculated for the same system or for a similar kind of material.
2.2. Meaning of Free Energy in Molecular System
In a thermodynamic system, Free energy is represented as equation (
17) as a measure of process progression since thermodynamic processes spontaneously proceed in the direction of decreasing system energy and increasing system disorder.
As mentioned above, the electron density in a molecular system can be regarded as a state quantity.
And the molecular formation process proceeds in the direction of decreasing total energy and increasing binding energy. Hence, the free energy in a molecular system can be defined as equation (
18).
where E is the total energy of the molecule, is the total electron density value of the molecule, which is the algebraic sum of the electron density values of each atom.
We defined the free energy of a molecule as
in the above expression and considered it as a measure of the direction of molecular formation and molecular optimization. The physical meaning of free energy can be called the difference between total energy and bound energy in both systems, and the value of free energy in the molecular system is the sum of the atomic region energies. This theory can be used to explain the phenomenon of the existence of a more stable state of a high-energy molecule for a molecule with the same chemical formula with potential well theory
| [17] | ZHAO Zhen-Min et al, Potential Energy Surface of Cytosine and Tunneling Between Its Normal and Trance-imino Tautomer, Commun. Theor. Phys. 46, (2006) 541-544. |
| [18] | Y. Cheng et al, Computational analysis of binding free energies between peptides and single-walled carbon nanotubes, Physica A 367, (2006) 293–304. |
| [19] | Lianzhong Deng et al, Optical Stark decelerator for molecules with a traveling potential well, PHYSICAL REVIEW A 95, 033409 (2017). |
| [20] | Jieli Qin and Lu Zhou, Unidirectional spin transport of a spin-orbit-coupled atomic matter wave using a moving Dirac δ-potential well, PHYSICAL REVIEW A 102, 013304 (2020). |
| [21] | HUNG Yong-Chang and WENG Gang, Solution of Wheeler-De Witt Equation, Potential Well and Tunnel Effect, Commun. Theor. Phys. 44, (2005) 757-761. |
[17-21]
.
2.3. Binding Energy of Water Molecule
Yet, despite its importance and ubiquitous ness in our lives, water remains uniquely unexplained compared to all the common chemicals we encounter, including many structural, dynamical and thermodynamic properties that underlie simple discernments which are required to accurately predict its behavior
| [12] | G. Zubay, Origins of Life on the Earth and in the Cosmos: On Earth and in the Cosmos, Academic Press, California, 2000. |
[12]
, This has led to a number of studies and controversies to predict and explain the physical and chemical behavior of water molecule and water molecule clusters.
The determination and solution of various improved models from empirical classical models for the behavior between water molecule clusters or water molecules has been investigated
| [13] | R. Aswani, J. C. Li, A new approach to pairwise potentials for water–water interactions, Journal of Molecular Liquids, 134 (2007) 120–128. |
| [14] | S. A. Volchek et al, Bond Energy in Nanostructured Water, International Journal of Nanoscience, Vol. 18, Nos. 3 & 4 (2019) 1940035. |
| [15] | Nancy Acelas et al, Structures, energies, and bonding in the water heptamer, J. Chem. Phys. 139, 044310 (2013). |
[13-15]
. These models have been explained on the basis of the “hydrogen bonding” of the dominant water molecules, and there have been many studies where molecular dynamics methods are applied to reproduce accurately the various properties determined by experiments.
In the study of the internal properties of water molecules, no accurate model determination and solution methods have been presented, especially the binding energy values of water molecules have not been accurately fitted.
The method of electron density can accurately fit the experimental values because it calculates the total binding energy, regardless of the kind, intensity, or number of interactions between atoms, and only the parameters that fit the species will be determined.
3. Calculation Method
To calculate the electron density values of the experimental molecules, a simulation for 100 molecules of 10 species was performed with a Gaussian 09w HF (6-31 G). The experimental values of the binding energy of the selected molecules was based on the literature
| [16] | Luo, Y R, Comprehensive Handbook of Chemical Bond Energies, CRC Press, 2007. |
[16]
.
4. Result and Discussion
In the figures of this paper, the unit for binding energy is (kcal/mol).
4.1. Correlation Between Binding Energy and Electron Density
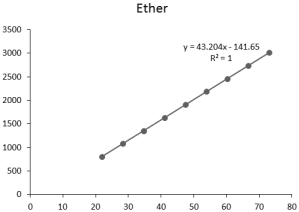
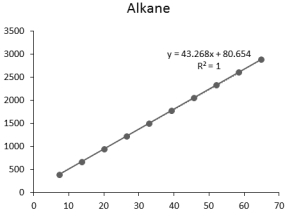
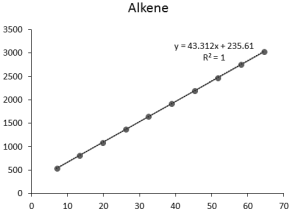
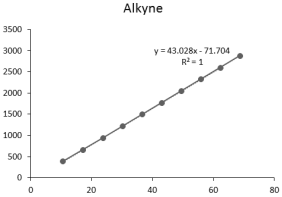
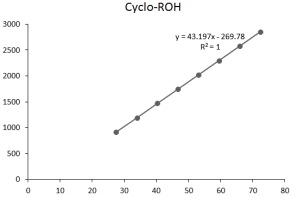
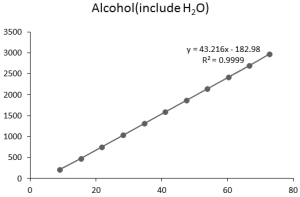
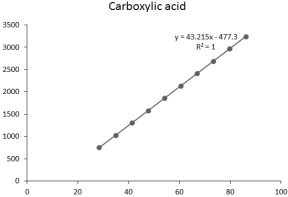
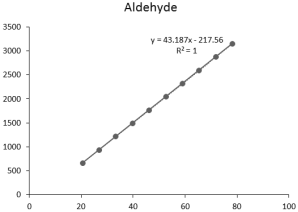
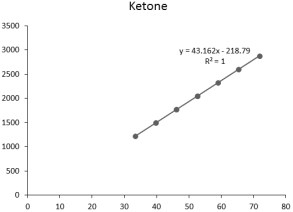
The correlation was investigated between the bond energy and the electron density since the value of the electron density in a molecule can reflect the binding energy of that molecule. The results showed a very significant correlation for each species in organic molecules, with a correlation value of more than 0.999.
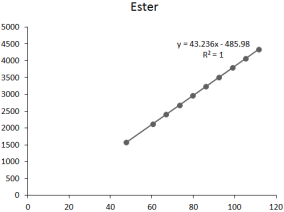
Figure 1. Correlation graph between the electron density and the binding energy for the species of molecule.
From the correlation between the total electron density and the binding energy of the molecule, the equation for calculating the binding energy using the electron density can be obtained.
(19)
The values of parameters for the some species are shown in the
Table 1.
Table 1. Values of parameters for the some species.
| x | y | R2 |
Alkane | 43.268 | 80.654 | 1 |
Alkene | 43.312 | 235.61 | 1 |
Alkyne | 43.028 | -71.704 | 1 |
Cyclo-ROH | 43.197 | -269.78 | 1 |
Alcohol | 43.216 | -182.98 | 0.9999 |
Carboxylic acid | 43.215 | -477.30 | 1 |
Aldehyde | 43.187 | -217.56 | 1 |
Ketone | 43.146 | -217.90 | 1 |
Ether | 43.204 | -141.65 | 1 |
Ester | 43.236 | -485.98 | 1 |
Interestingly, the first parameter, angular coefficient x, remains consistently around 43.2. This gives us the possibility of reducing the number of parameters.
4.2. Binding Energy of Water Molecule Calculated from Electron Density
Our method assumed that the total electron density value can reflect the total binding energy and accurately calculate the binding energy of water. And the results of the correlation study showed good result when matching alcohol species.
The electron density of each element in a water molecule is,
Table 2. Electron density of each element in a water molecule.
Element | O(1) | H(2) | H(3) |
Value of electron density | 8.28336 | 0.358377 | 0.358377 |
Of course, these values are the results of optimization in terms of the total energy of the molecule, rather than the free energy of the molecule.
However, there will be no or very little difference because the fraction of binding energy is small compared to the total energy and the structure with the optimum value of total energy is a stable molecule.
The parameter set for calculating the binding energy of water molecules from the electron density values is (43.216, -182.98) and result is 205.97. This value shows that the calculation accuracy of the binding energy of water is 94.2%.
4.3. Identification of the Theory on Optimization Direction Determination
We compared the total electron density before and after optimization for the organic molecule selectively by using the molecular calculation program (Chemoffice 2019, Gaussian09w). The comparison results show that the total electron density decreases in many cases after optimization, but there are also cases where the total electron density increases.
And the variation is not large. However, this suggests that the direction of molecular optimization cannot be viewed as a single factor effect alone.
For the validation of this theory, we have performed calculations for cytosine.
The total energy of cytosine reported in the literature is -393.952654 a.u
| [17] | ZHAO Zhen-Min et al, Potential Energy Surface of Cytosine and Tunneling Between Its Normal and Trance-imino Tautomer, Commun. Theor. Phys. 46, (2006) 541-544. |
[17]
. For this molecule, the optimum value for the HF (6-31 g) of Gaussian09w is -392.437362 a.u.
We tried to explain the difference between the experimental and simulated values from the calculated values before and after optimization by this theory.
Calculations of electron density, total energy and binding energy are given in
Table 3.
Table 3. Electron density, total energy and binding energy of cytosine.
Element | Electron density |
After | Before |
N(1) | 7.94294 | 7.88909 |
C(2) | 4.27363 | 4.27612 |
O(3) | 8.23077 | 8.18471 |
N(4) | 7.398 | 7.31756 |
C(5) | 4.50793 | 4.66891 |
N(6) | 7.33185 | 7.49147 |
C(7) | 5.29121 | 5.26517 |
C(8) | 4.70608 | 4.75068 |
H(9) | 0.286102 | 0.287748 |
H(10) | 0.298967 | 0.292888 |
H(11) | 0.322253 | 0.323109 |
H(12) | 0.432659 | 0.433492 |
H(13) | 0.421408 | 0.434942 |
Sum | 51.443799 | 51.615889 |
Total energy | -391.012705 a.u | -392.437462 a.u |
Binding energy | 2.059 a.u | 2.066 a.u |
The free energy change of the molecule is △Amol = 1.417757, and the actual total energy is -393.855219. The error to the actual value is 0.097435 and the calculation accuracy is 99.975%.
5. Conclusion
This work focuses on calculating the binding energy of a molecule based on the thermodynamic formulation. We have shown that the binding energy of a molecule can be calculated from the electron density by thermodynamic formalization.
The correlation between the electron density and the binding energy for 100 materials was studied for 10 species and the calculated expressions and parameters were determined from the interpolation polynomial. In this way, the accuracy of the calculation of the binding energy of water molecules by fitting the parameters of alcohol was 94.2%.
Also, a new theory on determination of molecular optimization direction is proposed through the analysis of the meaning of the free energy of a molecule in a molecular thermodynamic system. That is, molecular optimization are performed in the direction of decreasing the total energy of the molecule and of increasing the sum of the electron density, the binding energy.
This theory, with the potential well theory in quantum systems, could explain the phenomenon that high-energy molecules exist more stably. Since the proportion of binding energy is small compared to the total energy of a molecule, it is generally the same as the existing total energy minimization principle. However, it is different for some molecules and using this theory, more accurate calculations can be obtained. The proposed theory was validated by experimental and simulated values of cytosine and the results showed its effectiveness.
Abbreviations
WFT | Wave Function Theory |
DFT | Density Function Theory |
PCP | Pseudo Chemical Potential |
Acknowledgments
We acknowledge support data from the handbook of Luo, Y R – ' Comprehensive Handbook of Chemical Bond Energies'.
Thank Luo, Y R and ZHAO Zhen-Min et al presenting data of total and binding energy.
Author Contributions
Yong Myong Ri: Conceptualization, Investigation, Methodology, Supervision, Writing – original draft, Writing – review & editing
Tok Hui Ri: Supervision, Writing – review & editing, Methodology
Hak Sung Yun: Supervision, Writing – review & editing, Investigation
Guk Chol Kim: Investigation, Writing – review & editing, Methodology
Won Guk Ri: Investigation, Writing – review & editing, Methodology
Tong Il Kim: Writing – review & editing, Methodology, Validation
Data Availability Statement
The data supporting this study are available from the corresponding author on reasonable request.
Conflicts of Interest
The authors declare that they have no known competing financial interests or personal relationships that could have appeared to influence the work reported in this paper.
References
| [1] |
E. Schrödinger, An Undulatory Theory of the Mechanics of Atoms and Molecules. Phys. Rev. 28, 1049–1070 (1926).
|
| [2] |
Hartree, D. R. The Calculation of Atomic Structures. Rep. Prog. Phys. 11, 113-43 (1947).
|
| [3] |
Kohn, W. Nobel Lecture: Electronic Structure of Matter-Wave Functions and Density Functionals. Rev. Mod. Phys. 71, 1253–1266 (1999).
|
| [4] |
Miyajima, K., Yabushita, S., Knickelbein, M. B. & Nakajima, A. Stern-Gerlach Experiments of One-Dimensional Metal-Benzene Sandwich Clusters: Mn(C6H6)m (M) Al, Sc, Ti, and V). J. Am. Chem. Soc. 129, 8473–8480 (2007).
|
| [5] |
DiBenedetto, S. A. et al. Structure-Performance Correlations in Vapor Phase Deposited Self-Assembled Nanodielectrics for Organic Field-Effect Transistors. J. Am. Chem. Soc. 131, 11080–11090 (2009).
|
| [6] |
Prasad, V. K., Otero-de-la-Roza, A. & DiLabio, G. A. Atom-Centered Potentials with Dispersion-Corrected Minimal-Basis-Set Hartree-Fock: An Efficient and Accurate Computational Approach for Large Molecular Systems. J. Chem. Theory Comput. 14, 726–738 (2018).
|
| [7] |
Kohn, W. & Sham, L. J. Self-Consistent Equations Including Exchange and Correlation Effects. Phys. Rev. 140, A1133–A1138 (1965).
|
| [8] |
Slater, J. C. Quantum Theory of Matter, 2nd ed.; McGraw-Hill: New York, Chapter 16. 68).
|
| [9] |
T. I. Kim et al, A Novel Method for Calculation of Molecular Energies and Charge Distributions by Thermodynamic Formalization, Scientific Reports 9, (2019) 20264.
|
| [10] |
Jon Yung et al, Study on Calculation Method of Parameters in Molecular Calculation Model by Pseudo Chemical Potential Method, Science Research, (2023) 11(3): 43-50.
|
| [11] |
Jong Yong Ju et al, Theoretical Base of Pseudo Chemical Potential (PCP) Method, Science Research, (2022) 10(6): 137-143.
|
| [12] |
G. Zubay, Origins of Life on the Earth and in the Cosmos: On Earth and in the Cosmos, Academic Press, California, 2000.
|
| [13] |
R. Aswani, J. C. Li, A new approach to pairwise potentials for water–water interactions, Journal of Molecular Liquids, 134 (2007) 120–128.
|
| [14] |
S. A. Volchek et al, Bond Energy in Nanostructured Water, International Journal of Nanoscience, Vol. 18, Nos. 3 & 4 (2019) 1940035.
|
| [15] |
Nancy Acelas et al, Structures, energies, and bonding in the water heptamer, J. Chem. Phys. 139, 044310 (2013).
|
| [16] |
Luo, Y R, Comprehensive Handbook of Chemical Bond Energies, CRC Press, 2007.
|
| [17] |
ZHAO Zhen-Min et al, Potential Energy Surface of Cytosine and Tunneling Between Its Normal and Trance-imino Tautomer, Commun. Theor. Phys. 46, (2006) 541-544.
|
| [18] |
Y. Cheng et al, Computational analysis of binding free energies between peptides and single-walled carbon nanotubes, Physica A 367, (2006) 293–304.
|
| [19] |
Lianzhong Deng et al, Optical Stark decelerator for molecules with a traveling potential well, PHYSICAL REVIEW A 95, 033409 (2017).
|
| [20] |
Jieli Qin and Lu Zhou, Unidirectional spin transport of a spin-orbit-coupled atomic matter wave using a moving Dirac δ-potential well, PHYSICAL REVIEW A 102, 013304 (2020).
|
| [21] |
HUNG Yong-Chang and WENG Gang, Solution of Wheeler-De Witt Equation, Potential Well and Tunnel Effect, Commun. Theor. Phys. 44, (2005) 757-761.
|
Cite This Article
-
APA Style
Ri, Y. M., Ri, T. H., Yun, H. S., Kim, G. C., Ri, W. G., et al. (2026). A New Approach to Calculating the Binding Energy of Molecules from Electron Density by Thermodynamic Formalization. Science Discovery Physics, 1(1), 43-50. https://doi.org/10.11648/j.sdp.20260101.14
 Copy
|
 Download
ACS Style
Ri, Y. M.; Ri, T. H.; Yun, H. S.; Kim, G. C.; Ri, W. G., et al. A New Approach to Calculating the Binding Energy of Molecules from Electron Density by Thermodynamic Formalization. Sci. Discov. Phys. 2026, 1(1), 43-50. doi: 10.11648/j.sdp.20260101.14
Copy
|
Download
AMA Style
Ri YM, Ri TH, Yun HS, Kim GC, Ri WG, et al. A New Approach to Calculating the Binding Energy of Molecules from Electron Density by Thermodynamic Formalization. Sci Discov Phys. 2026;1(1):43-50. doi: 10.11648/j.sdp.20260101.14
Copy
|
Download
-
@article{10.11648/j.sdp.20260101.14,
  author = {Yong Myong Ri and Tok Hui Ri and Hak Sung Yun and Guk Chol Kim and Won Guk Ri and Tong Il Kim},
  title = {A New Approach to Calculating the Binding Energy of Molecules from Electron Density by Thermodynamic Formalization},
  journal = {Science Discovery Physics},
  volume = {1},
  number = {1},
  pages = {43-50},
  doi = {10.11648/j.sdp.20260101.14},
  url = {https://doi.org/10.11648/j.sdp.20260101.14},
  eprint = {https://article.sciencepublishinggroup.com/pdf/10.11648.j.sdp.20260101.14},
  abstract = {There are many approaches to molecular energy calculations, and there are still no methods to calculate the binding energy of a molecule accurately. Also, in considering molecules, all processes are actually performed at finite temperatures. And the process of molecular formation is a process of increasing the binding energy of a molecule, and if the molecule is considered as a thermodynamic system, it is also possible to think of the physical quantity that determines the direction of the spontaneous process. This paper propose a new approach to calculating the binding energy of a molecule by examining the relationship between the electron density and binding energy and about the thermodynamic formalization of molecule. From the calculated value by Gaussian09w (HF, 6-31g) simulation of 100 molecules of 10 species and the binding energy in some literatures, the correlation was derived and confirmed that two parameters are given for each species. It was found that the binding energy of a water molecule can be calculated, in particular. Also the molecular free energy is conceptualized where the value can be index of molecular formation and molecular optimization process. This theory, together with the potential well theory in quantum mechanics, can serve as a basis for explaining the phenomenon that for molecules with the same chemical structure formula, higher energy molecules exist more stably.},
 year = {2026}
}
Copy
|
Download
-
TY  - JOUR
T1  - A New Approach to Calculating the Binding Energy of Molecules from Electron Density by Thermodynamic Formalization
AU  - Yong Myong Ri
AU  - Tok Hui Ri
AU  - Hak Sung Yun
AU  - Guk Chol Kim
AU  - Won Guk Ri
AU  - Tong Il Kim
Y1  - 2026/02/25
PY  - 2026
N1  - https://doi.org/10.11648/j.sdp.20260101.14
DO  - 10.11648/j.sdp.20260101.14
T2  - Science Discovery Physics
JF  - Science Discovery Physics
JO  - Science Discovery Physics
SP  - 43
EP  - 50
PB  - Science Publishing Group
SN  - 3071-5458
UR  - https://doi.org/10.11648/j.sdp.20260101.14
AB  - There are many approaches to molecular energy calculations, and there are still no methods to calculate the binding energy of a molecule accurately. Also, in considering molecules, all processes are actually performed at finite temperatures. And the process of molecular formation is a process of increasing the binding energy of a molecule, and if the molecule is considered as a thermodynamic system, it is also possible to think of the physical quantity that determines the direction of the spontaneous process. This paper propose a new approach to calculating the binding energy of a molecule by examining the relationship between the electron density and binding energy and about the thermodynamic formalization of molecule. From the calculated value by Gaussian09w (HF, 6-31g) simulation of 100 molecules of 10 species and the binding energy in some literatures, the correlation was derived and confirmed that two parameters are given for each species. It was found that the binding energy of a water molecule can be calculated, in particular. Also the molecular free energy is conceptualized where the value can be index of molecular formation and molecular optimization process. This theory, together with the potential well theory in quantum mechanics, can serve as a basis for explaining the phenomenon that for molecules with the same chemical structure formula, higher energy molecules exist more stably.
VL  - 1
IS  - 1
ER  -
Copy
|
Download


 Share This Article
 Twitter
 Linked In
 Facebook